Amyotrophe Lateralsklerose
Zuletzt aktualisiert 18 Aug 2023
Erfüllt die redaktionelle Richtlinien
- HerunterladenHerunterladen
- Teilen
- Language
- Diskussion
- Audio-Version
- Zu bevorzugten Quellen bei Google hinzufügen
Amyotrophic lateral sclerosis (ALS) is a progressive disease that affects motor neurons, which are specialized nerve cells that control muscle movement. These nerve cells are found in the spinal cord and the brain. In ALS, motor neurons die (atrophy) over time, leading to muscle weakness, a loss of muscle mass, and an inability to control movement.
Beschreibung
Amyotrophic lateral sclerosis (ALS) is a progressive disease that affects motor neurons, which are specialized nerve cells that control muscle movement. These nerve cells are found in the spinal cord and the brain. In ALS, motor neurons die (atrophy) over time, leading to muscle weakness, a loss of muscle mass, and an inability to control movement.
There are many different types of ALS; these types are distinguished by their signs and symptoms and their genetic cause or lack of clear genetic association. Most people with ALS have a form of the condition that is described as sporadic, which means it occurs in people with no apparent history of the disorder in their family. People with sporadic ALS usually first develop features of the condition in their late fifties or early sixties. A small proportion of people with ALS, estimated at 5 to 10 percent, have a family history of ALS or a related condition called frontotemporal dementia (FTD), which is a progressive brain disorder that affects personality, behaviour, and language. The signs and symptoms of familial ALS typically first appear in one's late forties or early fifties. Rarely, people with familial ALS develop symptoms in childhood or their teenage years. These individuals have a rare form of the disorder known as juvenile ALS.
The first signs and symptoms of ALS may be so subtle that they are overlooked. The earliest symptoms include muscle twitching, cramping, stiffness, or weakness. Affected individuals may develop slurred speech (dysarthria) and, later, difficulty chewing or swallowing (dysphagia). Many people with ALS experience malnutrition because of reduced food intake due to dysphagia and an increase in their body's energy demands (metabolism) due to prolonged illness. Muscles become weaker as the disease progresses, and arms and legs begin to look thinner as muscle tissue atrophies. Individuals with ALS eventually lose muscle strength and the ability to walk. Affected individuals eventually become wheelchair-dependent and increasingly require help with personal care and other activities of daily living. Over time, muscle weakness causes affected individuals to lose the use of their hands and arms. Breathing becomes difficult because the muscles of the respiratory system weaken. Most people with ALS die from respiratory failure within 2 to 10 years after the signs and symptoms of ALS first appear; however, disease progression varies widely among affected individuals.
Approximately 20 percent of individuals with ALS also develop FTD. Changes in personality and behaviour may make it difficult for affected individuals to interact with others in a socially appropriate manner. Communication skills worsen as the disease progresses. It is unclear how the development of ALS and FTD are related. Individuals who develop both conditions are diagnosed as having ALS-FTD.
A rare form of ALS that often runs in families is known as ALS-parkinsonism-dementia complex (ALS-PDC). This disorder is characterised by the signs and symptoms of ALS, in addition to a pattern of movement abnormalities known as parkinsonism, and a progressive loss of intellectual function (dementia). Signs of parkinsonism include unusually slow movements (bradykinesia), stiffness, and tremors. Affected members of the same family can have different combinations of signs and symptoms.
Inheritance patterns
The source lists the following possible inheritance patterns: X-linked dominant; Not inherited; Autosomal dominant; Autosomal recessive. More than one pattern can apply in different cases.
Related genes
Other names
ALS
Amyotrophic lateral sclerosis with dementia
Charcot disease
Dementia with amyotrophic lateral sclerosis
Lou Gehrig disease
Motor neuron disease, amyotrophic lateral sclerosis
Related patient information and medicines
Patient information leaflet
Motoneuronerkrankung
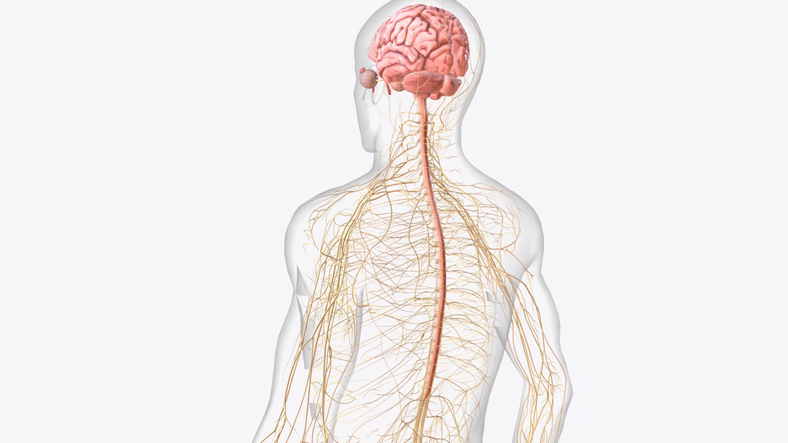
Motor neurone disease (MND) is a neurological condition causes a progressive weakness and wasting of many of the muscles in the body. This leaflet is mainly about amyotrophic lateral sclerosis (ALS), which is the most common type of motor neurone disease. Types of motor neurone disease The main types of MND are: Amyotrophic lateral sclerosis (ALS).
Patient information leaflet
Nervenschäden nach Epiduralinjektion
If you have a medical condition that interferes with blood supply (for example, diabetes ) or nerve function (for example, Multiple Sklerose ), this can make damage more likely or make it more difficult to know exactly what has caused the nerve damage. Der problems with the nerves may also be due to a health condition you already have, such as a.
Patient information leaflet
Das Immunsystem
This leaflet gives a brief overview of the Immunsystem system and how it works. A person with a weak immune system might have difficulty fighting off infections. In autoimmune disea... Autoimmune diseases can attack any part of the body - eg, type 1 diabetes (pancreas), Multiple Sklerose (brain and nerves), systemic lupus erythematosus (skin and.
Patient information leaflet
COVID-19
See the long COVID leaflet for. How long are you contagious with COVID. Most people will no longer be contagious at five days after developing symptoms of COVID. Conditions causing weakene... Medical treatments causing weakened immune systems (like steroid medicines , antibody treatments, or chemotherapy ). Rare conditions affecting the brain and.
Patient information leaflet
Blasenpemphigoid
Blasenpemphigoid ist a skin disease that can cause a rash, itching and blisters. It mainly affects people aged over 70. Treatment usually works well to control symptoms. Treatment is usually with steroid creams or medicines, but sometimes other medicines may be used. The condition tends to go away after 3-6 years and then treatment can be.
Patient information leaflet
Trigeminusneuralgie
Trigeminusneuralgie is defined as nerve pain (neuralgia) involving one or more of the branches of the trigeminal nerves. The trigeminal nerve carries sensation from your face to your brain. Trigeminal neuralgia is a condition which causes sudden, severe facial pain. ... Therefore, a typical person with trigeminal neuralgia is an older person, has.
Weiterführende Lektüre und Referenzen
- Amyotrophe Lateralsklerose; MedlinePlus, National Library of Medicine.
- Andersen PM, Al-Chalabi A. Clinical genetics of amyotrophic lateral sclerosis: what do we really know? Nat Rev Neurol. 2011 Oct 11;7(11):603-15. doi: 10.1038/nrneurol.2011.150.
- Andersen PM. Amyotrophic lateral sclerosis associated with mutations in the CuZn superoxide dismutase gene. Curr Neurol Neurosci Rep. 2006 Jan;6(1):37-46. doi: 10.1007/s11910-996-0008-9.
- Chio A, Borghero G, Restagno G, Mora G, Drepper C, Traynor BJ, Sendtner M, Brunetti M, Ossola I, Calvo A, Pugliatti M, Sotgiu MA, Murru MR, Marrosu MG, Marrosu F, Marinou K, Mandrioli J, Sola P, Caponnetto C, Mancardi G, Mandich P, La Bella V, Spataro R, Conte A, Monsurro MR, Tedeschi G, Pisano F, Bartolomei I, Salvi F, Lauria Pinter G, Simone I, Logroscino G, Gambardella A, Quattrone A, Lunetta C, Volanti P, Zollino M, Penco S, Battistini S; ITALSGEN consortium; Renton AE, Majounie E, Abramzon Y, Conforti FL, Giannini F, Corbo M, Sabatelli M. Clinical characteristics of patients with familial amyotrophic lateral sclerosis carrying the pathogenic GGGGCC hexanucleotide repeat expansion of C9ORF72. Brain. 2012 Mar;135(Pt 3):784-93. doi: 10.1093/brain/awr366.
- Farg MA, Sundaramoorthy V, Sultana JM, Yang S, Atkinson RA, Levina V, Halloran MA, Gleeson PA, Blair IP, Soo KY, King AE, Atkin JD. C9ORF72, implicated in amytrophic lateral sclerosis and frontotemporal dementia, regulates endosomal trafficking. Hum Mol Genet. 2014 Jul 1;23(13):3579-95. doi: 10.1093/hmg/ddu068. Epub 2014 Feb 18.
- Fecto F, Siddique T. Making connections: pathology and genetics link amyotrophic lateral sclerosis with frontotemporal lobe dementia. J Mol Neurosci. 2011 Nov;45(3):663-75. doi: 10.1007/s12031-011-9637-9. Epub 2011 Sep 7.
- Ferraiuolo L, Kirby J, Grierson AJ, Sendtner M, Shaw PJ. Molecular pathways of motor neuron injury in amyotrophic lateral sclerosis. Nat Rev Neurol. 2011 Nov;7(11):616-30. doi: 10.1038/nrneurol.2011.152.
- Kiernan MC, Vucic S, Cheah BC, Turner MR, Eisen A, Hardiman O, Burrell JR, Zoing MC. Amyotrophic lateral sclerosis. Lancet. 2011 Mar 12;377(9769):942-55. doi: 10.1016/S0140-6736(10)61156-7. Epub 2011 Feb 4.
- Lattante S, Ciura S, Rouleau GA, Kabashi E. Defining the genetic connection linking amyotrophic lateral sclerosis (ALS) with frontotemporal dementia (FTD). Trends Genet. 2015 May;31(5):263-73. doi: 10.1016/j.tig.2015.03.005. Epub 2015 Apr 10.
- Ling SC, Polymenidou M, Cleveland DW. Converging mechanisms in ALS and FTD: disrupted RNA and protein homeostasis. Neuron. 2013 Aug 7;79(3):416-38. doi: 10.1016/j.neuron.2013.07.033.
- Marangi G, Traynor BJ. Genetic causes of amyotrophic lateral sclerosis: new genetic analysis methodologies entailing new opportunities and challenges. Brain Res. 2015 May 14;1607:75-93. doi: 10.1016/j.brainres.2014.10.009. Epub 2014 Oct 12.
- McGeer PL, Steele JC. The ALS/PDC syndrome of Guam: potential biomarkers for an enigmatic disorder. Prog Neurobiol. 2011 Dec;95(4):663-9. doi: 10.1016/j.pneurobio.2011.04.001. Epub 2011 Apr 20.
- National Institute of Neurological Disorders and Stroke
- Paganoni S, Macklin EA, Lee A, Murphy A, Chang J, Zipf A, Cudkowicz M, Atassi N. Diagnostic timelines and delays in diagnosing amyotrophic lateral sclerosis (ALS). Amyotroph Lateral Scler Frontotemporal Degener. 2014 Sep;15(5-6):453-6. doi: 10.3109/21678421.2014.903974. Epub 2014 Jul 1.
- Renton AE, Chio A, Traynor BJ. State of play in amyotrophic lateral sclerosis genetics. Nat Neurosci. 2014 Jan;17(1):17-23. doi: 10.1038/nn.3584. Epub 2013 Dec 26.
- Siddique N, Siddique T. Amyotrophic Lateral Sclerosis Overview. 2001 Mar 23 [updated 2023 Sep 28]. In: Adam MP, Bick S, Mirzaa GM, Wallace SE, Amemiya A, editors. GeneReviews(R) [Internet]. Seattle (WA): University of Washington, Seattle; 1993-2026. Available from http://www.ncbi.nlm.nih.gov/books/NBK1450/
- Vance C, Rogelj B, Hortobagyi T, De Vos KJ, Nishimura AL, Sreedharan J, Hu X, Smith B, Ruddy D, Wright P, Ganesalingam J, Williams KL, Tripathi V, Al-Saraj S, Al-Chalabi A, Leigh PN, Blair IP, Nicholson G, de Belleroche J, Gallo JM, Miller CC, Shaw CE. Mutations in FUS, an RNA processing protein, cause familial amyotrophic lateral sclerosis type 6. Science. 2009 Feb 27;323(5918):1208-1211. doi: 10.1126/science.1165942.
Artikelverlauf
Artikel auch verfügbar in Englisch, Deutsch, Spanisch, Französisch, Italienisch, Portugiesisch, Hindi, Hebräisch, Arabisch, und Schwedisch.
18 Aug 2023 | Neueste Version
1 Feb 2016 | Ursprünglich veröffentlicht
Nehmen Sie an der Diskussion teil
Teilen Sie Ihre Gedanken oder Erfahrungen mit anderen in unserem Patientengemeinschaften.

Fragen, teilen, verbinden.
Durchsuchen Sie Diskussionen, stellen Sie Fragen und teilen Sie Erfahrungen zu Hunderten von Gesundheitsthemen.
Abonnieren Sie den Patienten-Newsletter
Ihre wöchentliche Dosis klarer, vertrauenswürdiger Gesundheitsberatung - geschrieben, um Ihnen zu helfen, sich informiert, selbstbewusst und in Kontrolle zu fühlen.
Durch das Abonnieren akzeptieren Sie unsere Datenschutzrichtlinie. Sie können sich jederzeit abmelden. Wir verkaufen Ihre Daten niemals.